Research
A connected view of my research
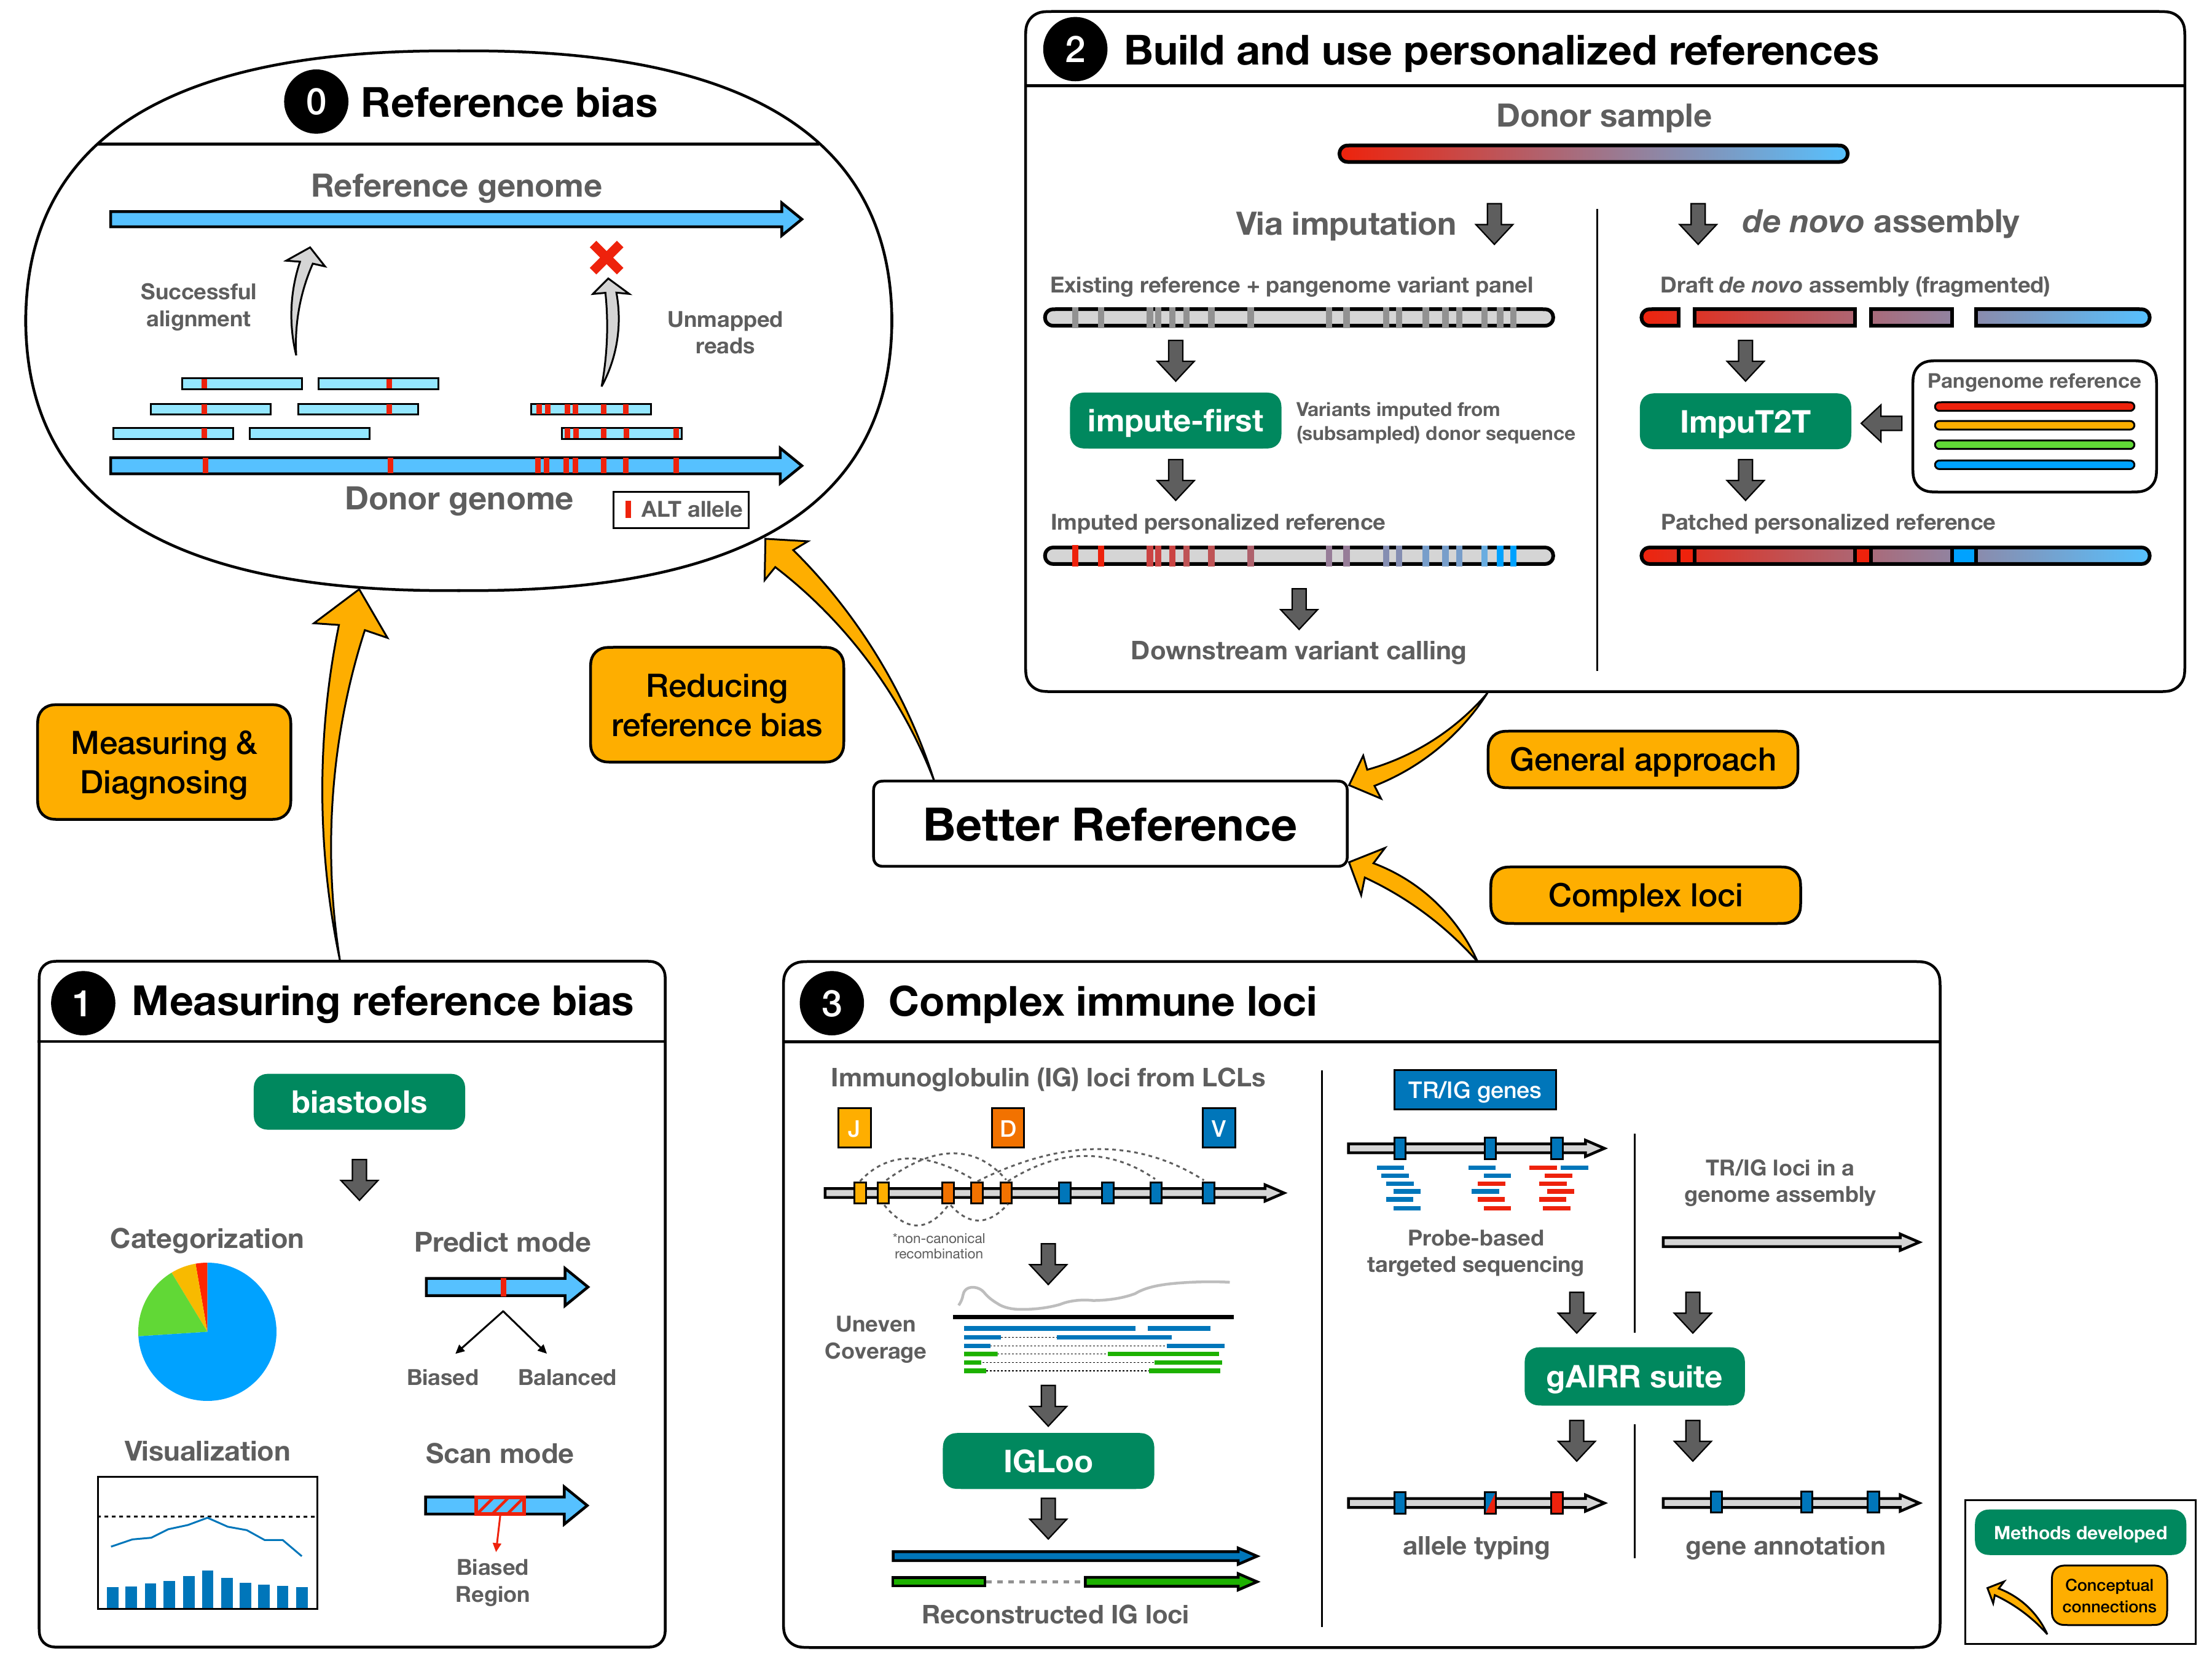
My work connects the measurement of reference bias with the development of personalized references and the reconstruction and profiling of complex immune loci. Together, these approaches aim to build better references for genomic analysis.
Select a project block to explore the work below. Select a numbered circle to enlarge its diagram.
0 · Background: reference bias
Reference bias occurs when reads carrying alternative (ALT) alleles fail to align correctly. The resulting alignment favors the reference genome’s genotype, which can affect downstream analysis. My work addresses both how to measure reference bias and how better reference genomes can reduce the bias.
1 · Measuring reference bias
biastools
Many alignment methods aim to reduce reference bias, but evaluating how much bias remains can be challenging. Reference bias can manifest in different ways, for example, a variant site may repel reads carrying the ALT allele, or it may attract reads from elsewhere in the genome, leading to misleading results. We developed biastools, a framework to measure, categorize, and visualize reference bias. Its analysis connects individual alleles and biased genomic regions to the overall behavior of an alignment workflow.
2 · Building and using personalized references
Personalized references bring the reference sequence closer to the donor genome. The diagram connects two routes: imputation before alignment, and patching a draft assembly with a pangenome reference.
impute-first
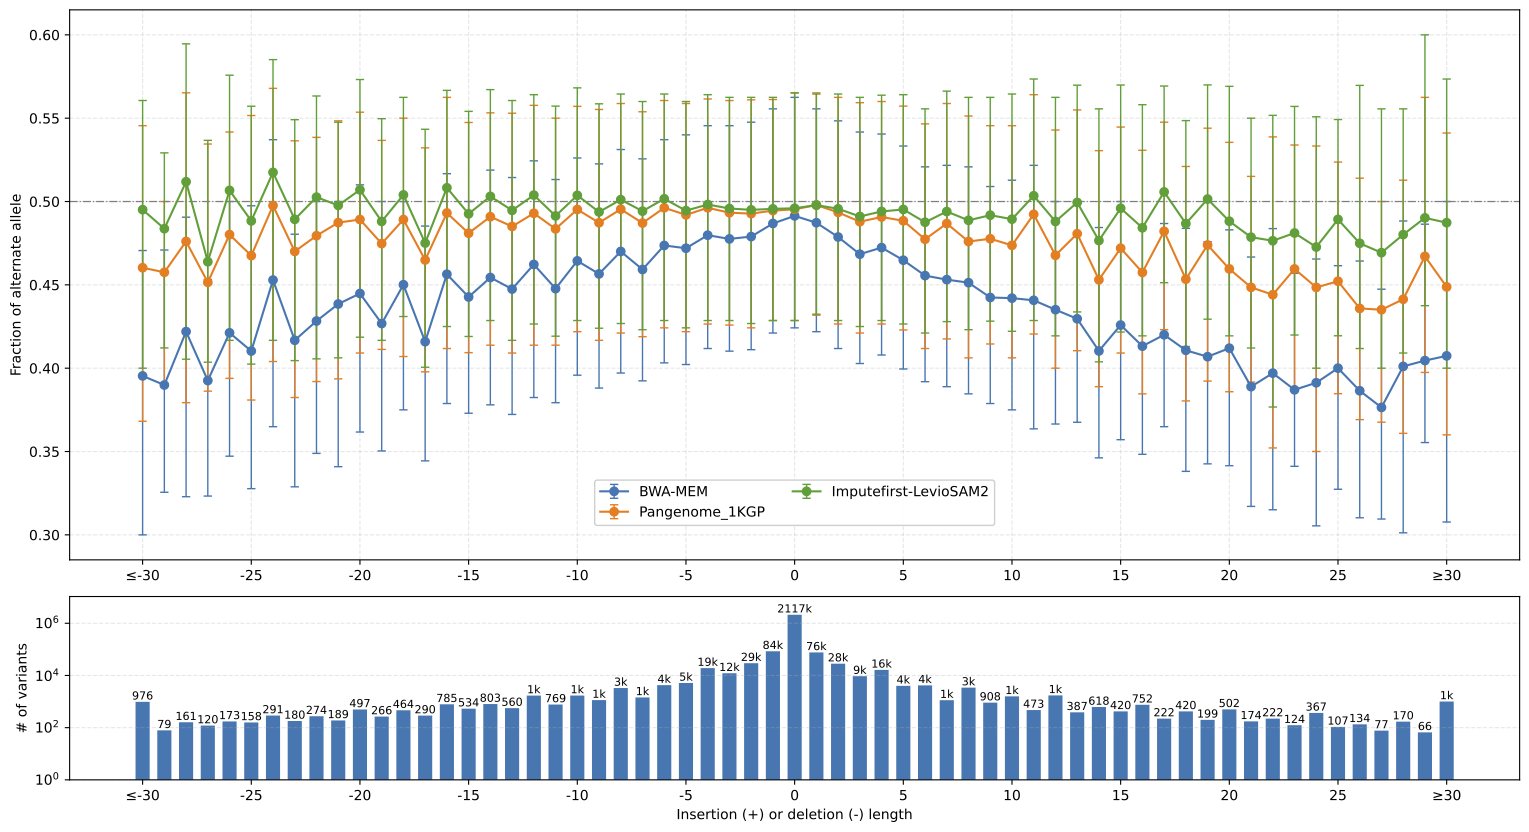
The impute-first project first imputes the donor’s genomic variants using a subsampled read alignment. This approach enables the creation of a personalized reference genome prior to the main sequence alignment step, which reduces reference bias and improves the accuracy of downstream variant calling.
I developed the downstream workflow using LevioSAM2 to lift alignments from the personalized genome back to a standard reference such as GRCh38 or T2T-CHM13. This approach enables the advantages of personalized references while maintaining compatibility with standard genomic coordinates, offering a linear-space alternative to graph-based alignment methods.
ImpuT2T
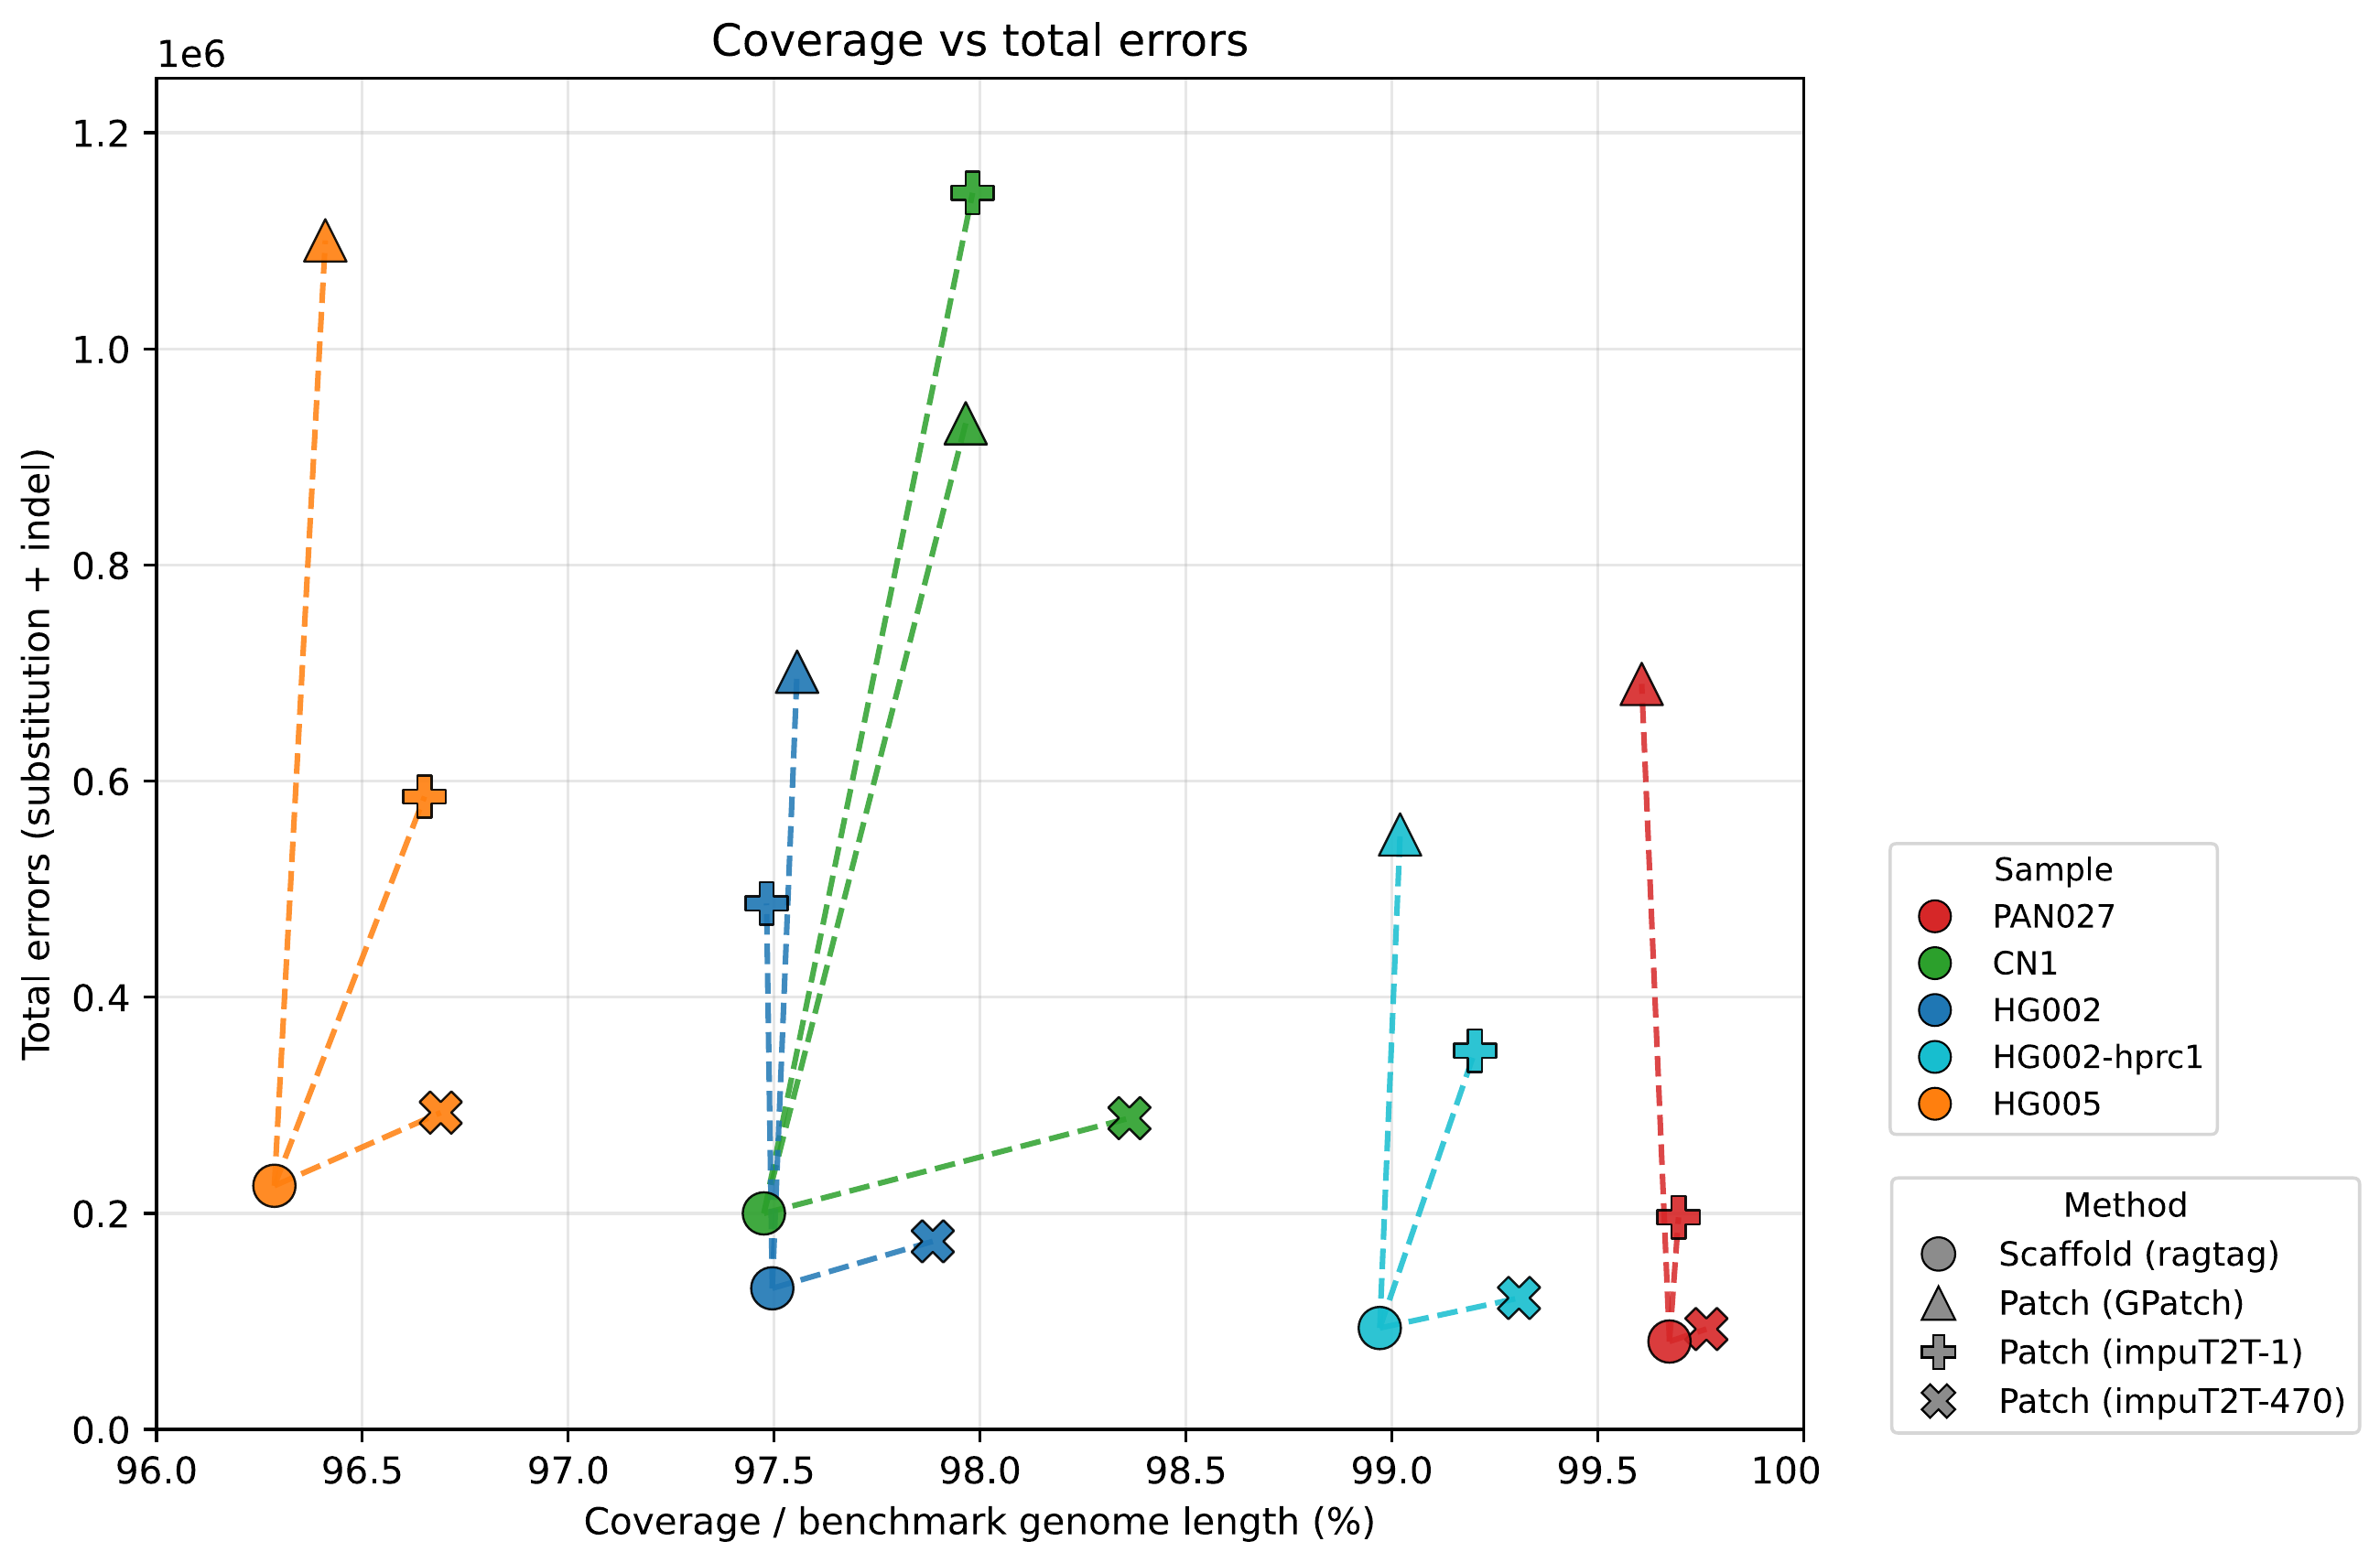
ImpuT2T utilizes a pangenome (HPRC2) to scaffold and patch draft genome assemblies, selecting the reference haplotype with the highest sequence identity to fill each gap. We compared standard reference-based scaffolding with RagTag, single-reference patching using GPatch, ImpuT2T with a single reference, and ImpuT2T using the full pangenome reference set. Among all methods, ImpuT2T with the pangenome recovered the largest portion of the benchmark genome while introducing the fewest errors.
3 · Resolving complex immune loci
Immunoglobulin (IG) and T cell receptor (TR) loci are genetically complex regions central to adaptive immunity. My work combines reconstruction of IG loci with germline gene profiling through targeted sequencing and genome assemblies.
IGLoo
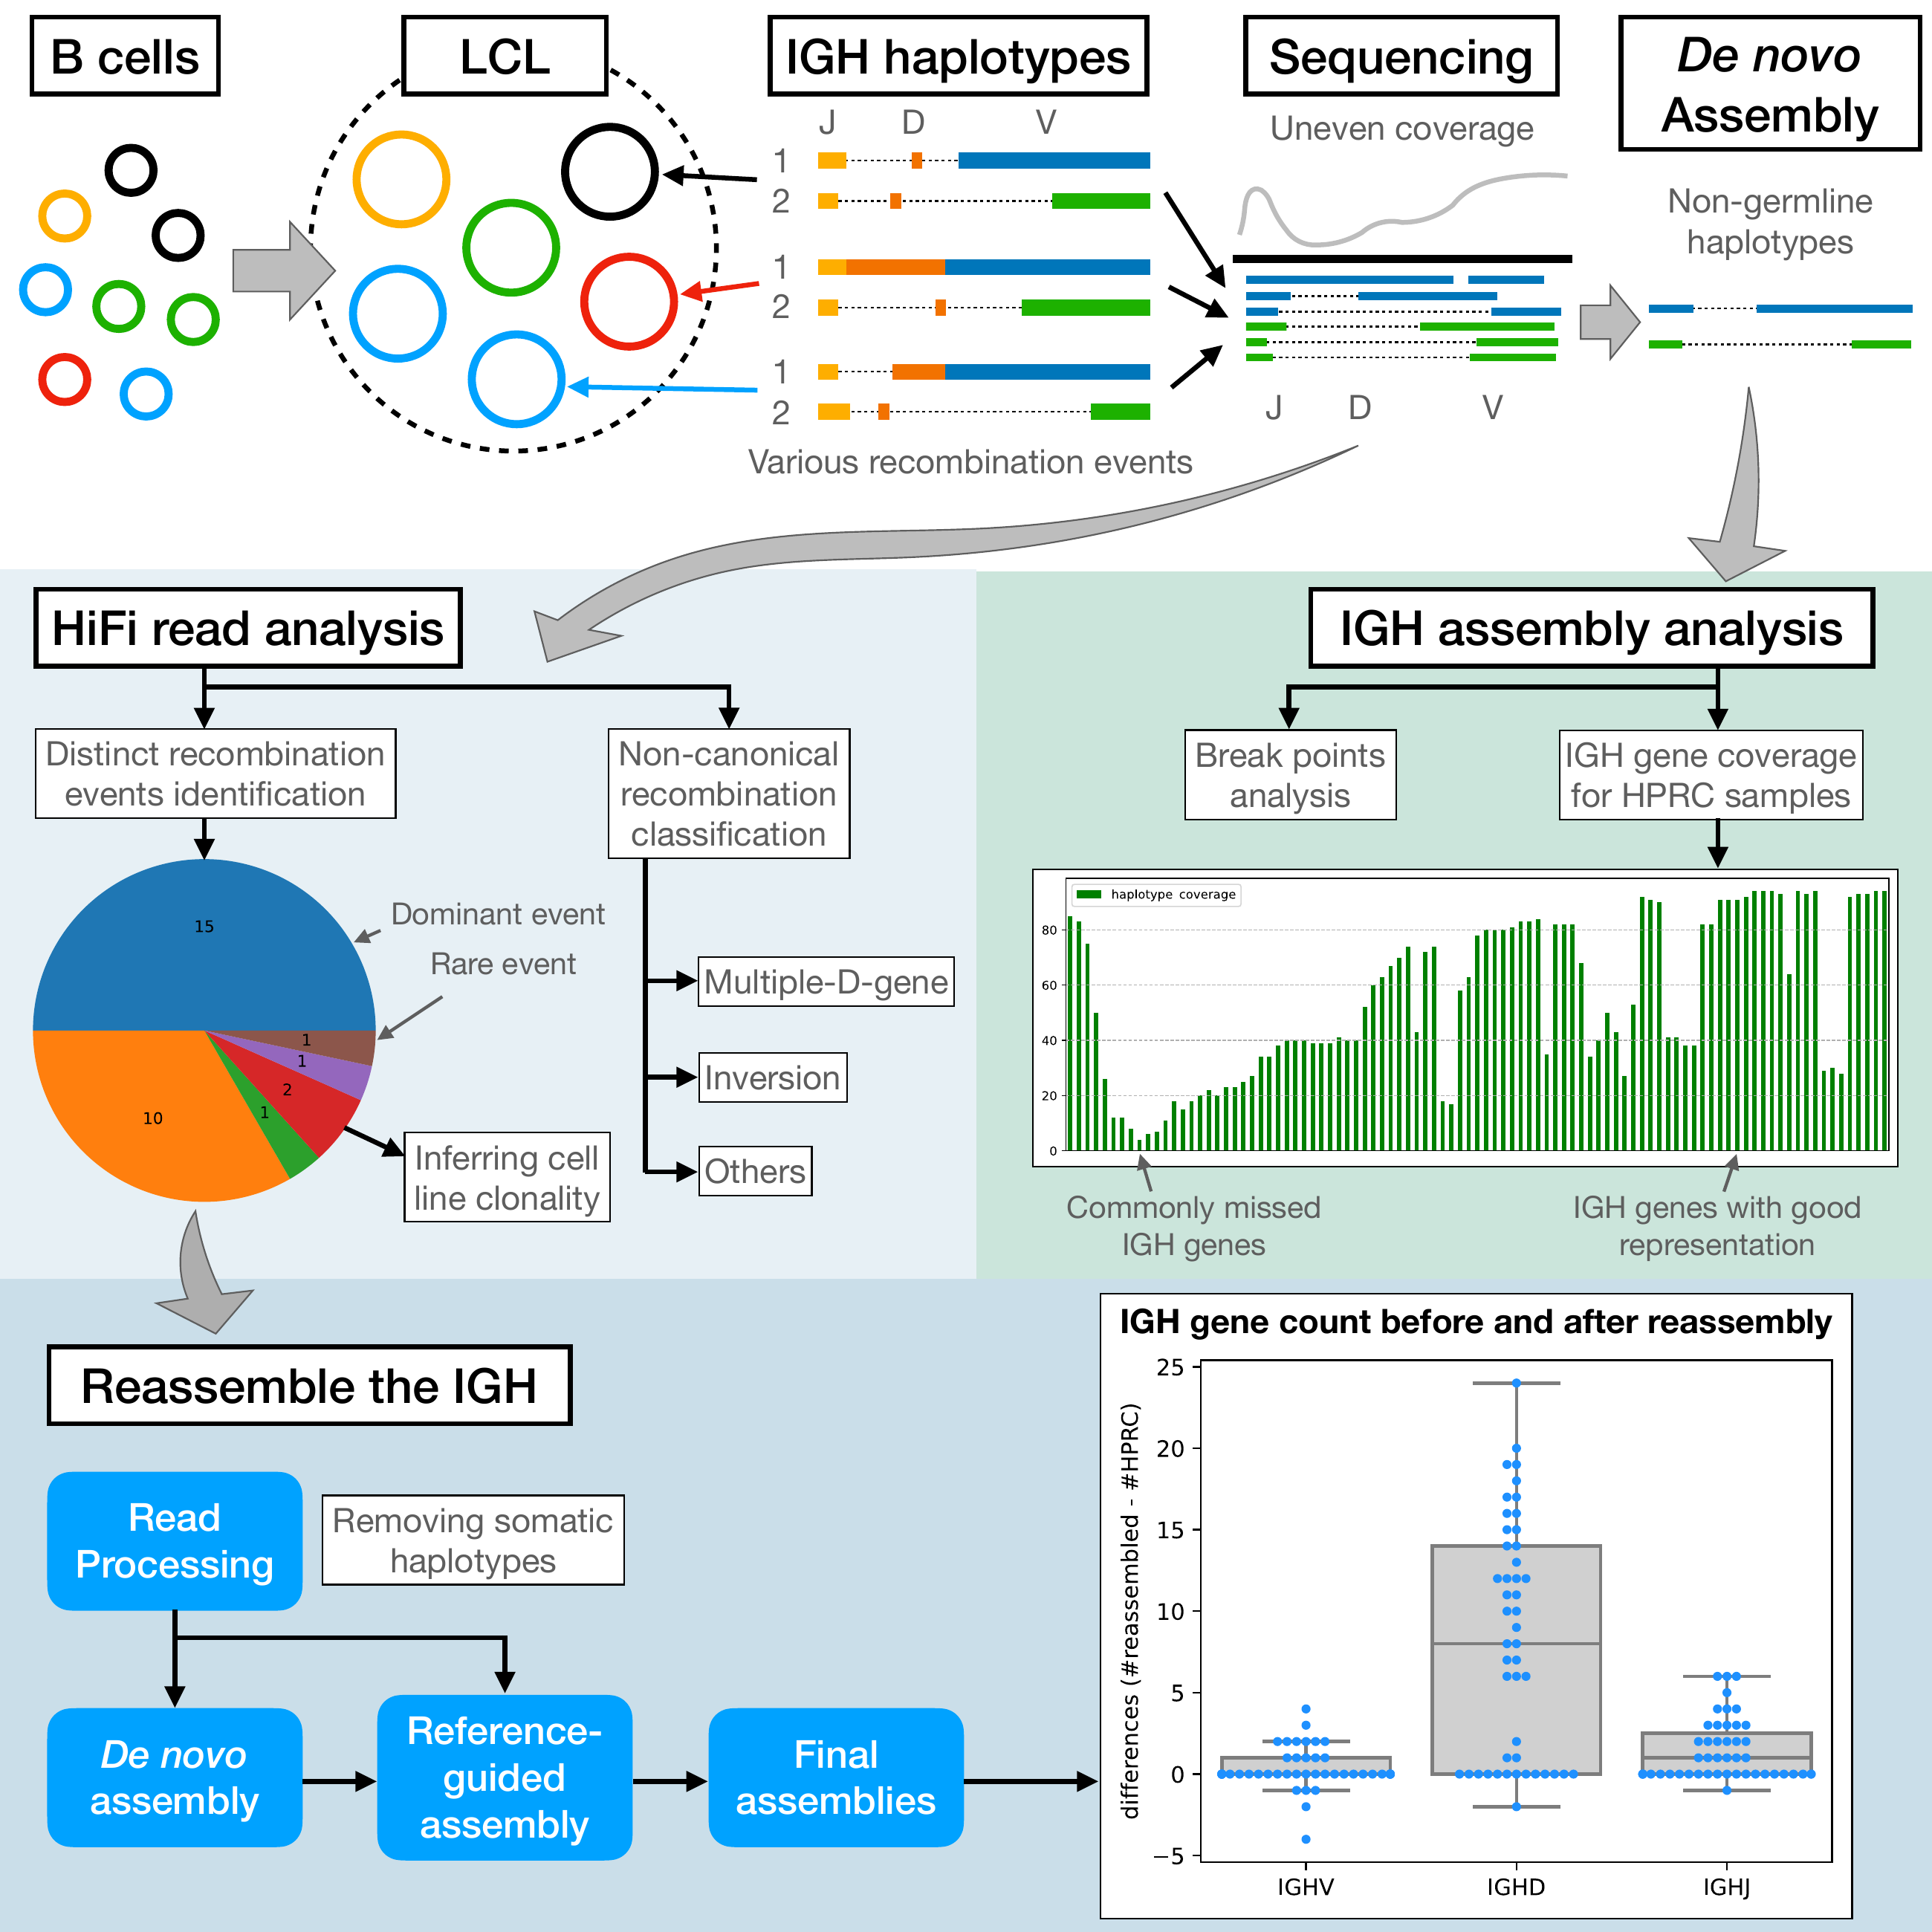
High-quality human genome assemblies derived from lymphoblastoid cell lines (LCLs) contribute to reference genomes and pangenomes. Their IG loci, however, can contain a mixture of germline and somatically recombined haplotypes, complicating genotyping and assembly.
We developed IGLoo to analyze V(D)J recombination events in LCL sequencing data and use that information to reassemble the immunoglobulin heavy chain (IGH) locus. Compared with the original HPRC-v1 assemblies, the reassembled loci contain more IG genes and have a lower overall switching error rate.
gAIRR-suite
The adaptive immune receptor repertoire (AIRR) is encoded by TR and IG genes. Profiling the germline genes that encode this repertoire (gAIRR) is important for understanding adaptive immune responses, but their genetic complexity presents substantial challenges.
We developed gAIRR-suite to profile human TR and IG genes using publicly available personal phased assemblies and capture-based targeted sequencing of genomic DNA. As shown in the diagram, these complementary inputs support gene annotation and allele typing.